Institute of Technology, Resource and Energy-efficient Engineering (TREE)
Multiscale Molecular Modelling Center
„Creating and using models have a long history in scientific research and education. Computational chemistry is an exciting field that requires cross-disciplinary knowledge to build realistic and trustworthy models. Because of this, one always has the opportunity to learn something new, apply ideas from one field to others, and create new knowledge at incredible resolution levels.”
Dr. Karl N. Kirschner
Research Overview
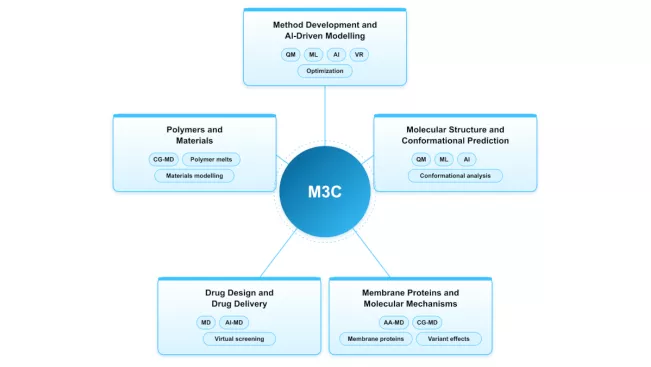
Overview of the center's topics and methods.
Method Development & AI Data-Driven Modelling
The central pillar of our research is the development and refinement of physics-based and data-driven methods. In force-field optimization, we focus on formulating tailored loss functions, tuning weighting parameters, and leveraging multi-objective optimization algorithms that preserve physical validity. Concurrently, we harness modern machine learning (ML) to explore complex parameter landscapes (utilizing advanced techniques like quality-diversity optimization), train surrogate models for molecular dynamics, and predict high-dimensional potential energy surfaces. We also apply ML to evaluate molecular similarity, generating novel hypotheses for experimental validation (e.g., drug repurposing). To maximize computational efficiency, we integrate data-driven models directly with quantum mechanical calculations to build hybrid ML → QM workflows.
Topics & Projects
- Force field & method development: loss functions, weighting factors, multi-objective optimization
- Correlation functions and optimization of the potential energy surface (PES)
- Method development and application of partial atomic charges, including the modeling of anisotropic electrostatics (σ -hole, electron lone-pairs)
- Multi-objective and quality-diversity optimization of the Lennard-Jones parameter landscape
- ML→QM workflow methodology
- AI-driven modelling and generative methods
- Visualization & teaching: virtual reality in teaching, proteins/MD in public outreach and as a video game
- Education: SciPro course; boiling points using ML with Jupyter Notebook
Molecular Structure & Conformational Prediction
Our group also predicts the molecular structures and conformations of small molecules. We explore conformational space by systematically rotating torsion angles, performing QM optimizations, and by integrating modern generative ML methods. For example, we embed QM calculations directly into the conformational search — enabling end-to-end ML → QM workflows that are applied to complex biopolymers (e.g., lignin). Our research investigates fundamental conformational phenomena, including ring-puckering dynamics in small cycles (e.g., cyclopentane), conformer ensembles for flexible hydrocarbons (e.g., octane), and σ-hole interactions between halogenated compounds and various organic molecules.
Topics & Projects
- AI-driven structure & conformational prediction of small molecules: benchmarking modern generative AI methods
- Extension of the conformational dataset (QM) for ML training
- QM in conformational generation for lignin
- ML→QM workflow methodology
- Cyclopentane ring puckering
- Nonane QM conformers
- Applications of force field development: 1-bromobutane, 2-bromobutane and extension to Cl, F, Br, I
- Br-substituted benzene – water interactions (QM)
Membrane Proteins & Molecular Mechanisms
Molecule Dynamics Modelling
A major focus of our research is investigating membrane proteins and understanding their underlying molecular mechanisms. Using all-atom and coarse-grained molecular dynamics (AA-MD and CG-MD), we analyze the structure, dynamics, and function of membrane transport and receptor systems — including the nicotinic acetylcholine receptor (nAChR) across distinct functional states, the sodium-proton exchanger NHE1, and organic cation (OCT) transporters. Our work also encompasses characterizing potential substrates for monocarboxylate transporter 1 (MCT1) to elucidate their role in congenital metabolic disorders.
Variant Effect Prediction & Cross-Species Modeling
Beyond direct molecular simulations, we develop ML models for variant effect prediction (VEP). We examine missense variant functional consequences across species boundaries (e.g., in ENaC and nAChR) and extend curated human VEP datasets by integrating data from mouse and rat models. This cross-species perspective helps bridge sparse-data gaps and improves predictive power.
Topics & Projects
- Membrane proteins & molecular mechanisms: nAChR (AA-MD, CG-MD; resting state 7KOO, open state 7KOX), NHE1 (AA-MD), OCT transporters (AA-MD)
- Variant effects & transport proteins: variant effect prediction across species boundaries (ENaC, nAChR)
- Cross-species variant effect prediction (VEP): extension of human VEP datasets with mouse and rat data
- Characterization of potential MCT1 substrates in connection with congenital metabolic disorders
Drug Design & Drug Delivery
Our group researches computer-aided strategies towards drug design and drug delivery. One focus is the analysis of structure-activity relationships and virtual screening — leveraging our custom MolFuSE framework. In drug delivery, we investigate carrier-mediated transport and controlled release mechanics, with a special emphasis on modeling cyclodextrin complexes. We study these systems using hybrid ML-driven molecular dynamics simulations (ML-MD) to bridge theoretical predictions with empirical data.
Topics & Projects
- Drug design & virtual screening: analysis of structure-activity relationships and virtual screening using the MolFuSE method
- Drug delivery & cyclodextrins: concepts of drug design and delivery as well as modelling of cyclodextrins using AI-driven molecular dynamics (AI-MD)
Polymers & Materials
A further research area is dedicated to the modelling of polymers and materials. The focus is on the investigation of polymer melts, in particular of high-density polyethylene (HDPE) for extrusion blow moulding, which is simulated using coarse-grained molecular dynamics (CG-MD). These efforts are extended to additives and to polypropylene (PP). In addition, modified membranes are analytically characterized and structurally interpreted.
Topics & Projects
- Polymers & materials: polymer melts of high-density polyethylene (HDPE) for extrusion blow moulding (CG-MD) with extension to additives and polypropylene (PP)
- Analytical characterization of modified membranes: structural interpretation of modified membrane systems
Labs
The calculations are performed on the high-performance computing cluster at Bonn-Rhein-Sieg University of Applied Sciences. Opportunities for efficient parallelization, combined with state-of-the-art simulation tools, enable highly detailed calculations.
Teaching
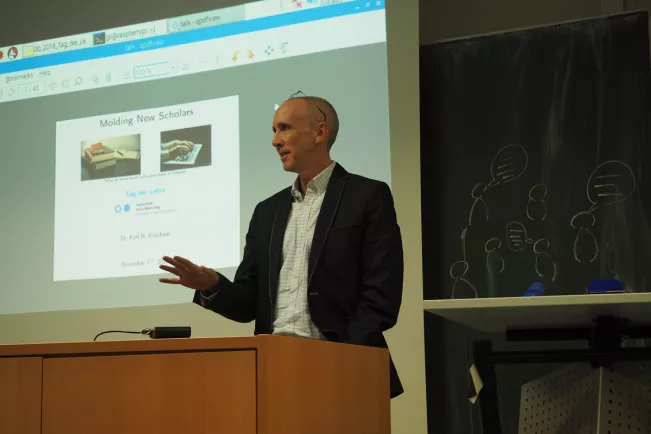
As part of the UMMBAS research project, the group is working on the development of a collaborative VR environment to enable efficient collaboration on topics in the field of molecular modeling across geographical boundaries. Once the collaboration concepts under development are established, they are expected to be applicable in teaching as well. In particular, they could be used in teaching modules offered across departments or universities.
Alumni
Partners
- Edward E. Whitacre Jr. College of Engineering, Texas Tech University (Prof. Dr. Roland Faller)
- Institut für Integrierte Naturwissenschaften, Universität Koblenz (Prof. Dr. Wolfgang Imhof)
- Institut für Prozess- und Verfahrenstechnik, Technische Universität Berlin (Prof. Dr. Jadran Vrabec)
- Department of Chemistry, Furman University, South Carolina (Prof. Dr. George Shields)
- Division of Hematology, Ohio State University, Comprehensive Cancer Center, Columbus, OH USA (Prof. Drs. John Byrd and Karilyn Larkin)
Selected Publications
Arbeitsgruppe Computational Chemistry
2023
Alexander Hagg, Karl N. Kirschner:
Open-Source Machine Learning in Computational Chemistry.
PDF Download (CC BY-NC-ND 4.0) doi:10.1021/acs.jcim.3c00643 PMID urn:nbn:de:hbz:1044-opus-74406
BibTeX | RIS
Max Müller, Alexander Hagg, Robin Strickstrock, Marco Hülsmann, Alexander Asteroth, Karl N. Kirschner, Dirk Reith:
Determining Lennard-Jones Parameters Using Multiscale Target Data through Presampling-Enhanced, Surrogate-Assisted Global Optimization.
doi:10.1021/acs.jcim.2c01231 PMID
BibTeX | RIS
JinHeng Lin, Sean M. Gettings, Khaoula Talbi, Rainer Schreiber, Michael J. Taggart, Matthias Preller, Karl Kunzelmann, Mike Althaus, Michael A. Gray:
Pharmacological inhibitors of the cystic fibrosis transmembrane conductance regulator exert off-target effects on epithelial cation channels.
PDF Download (CC BY 4.0) doi:10.1007/s00424-022-02758-9 PMID urn:nbn:de:hbz:1044-opus-64572
BibTeX | RIS
2022
Dirk Grommes, Martin R. Schenk, Olaf Bruch, Dirk Reith:
Initial Crystallization Effects in Coarse-Grained Polyethylene Systems After Uni- and Biaxial Stretching in Blow-Molding Cooling Scenarios.
PDF Download (CC BY 4.0) doi:10.3390/polym14235144 PMID urn:nbn:de:hbz:1044-opus-65219
BibTeX | RIS
Walter Fiedler, Fabian Freisleben, Jasmin Wellbrock, Karl N. Kirschner:
Mebendazole's Conformational Space and Its Predicted Binding to Human Heat-Shock Protein 90.
doi:10.1021/acs.jcim.2c00290 PMID
BibTeX | RIS
Robin Strickstrock, Marco Hülsmann, Dirk Reith, Karl N. Kirschner:
Optimizing Lennard-Jones parameters by coupling single molecule and ensemble target data.
doi:10.1016/j.cpc.2022.108285
BibTeX | RIS
2020
Meera C. Viswanathan, William Schmidt, Peter Franz, Michael J. Rynkiewicz, Christopher S. Newhard, Aditi Madan, William Lehman, Douglas M. Swank, Matthias Preller, Anthony Cammarato:
A role for actin flexibility in thin filament-mediated contractile regulation and myopathy.
doi:10.1038/s41467-020-15922-5 PMID
BibTeX | RIS
Wiebke Ewert, Peter Franz, Georgios Tsiavaliaris, Matthias Preller:
Structural and Computational Insights into a Blebbistatin-Bound Myosin•ADP Complex with Characteristics of an ADP-Release Conformation along the Two-Step Myosin Power Stoke.
PDF Download (CC BY 4.0) doi:10.3390/ijms21197417 urn:nbn:de:hbz:1044-opus-50854 PMID
BibTeX | RIS
Karl N. Kirschner, Dirk Reith, Wolfgang Heiden:
The performance of Dunning, Jensen, and Karlsruhe basis sets on computing relative energies and geometries.
doi:10.1080/1539445x.2020.1714656
BibTeX | RIS
Martin R. Schenk, Thorsten Köddermann, Karl N. Kirschner, Sandra Knauer, Dirk Reith:
Molecular Dynamics in the Energy Sector: Experiment and Modeling of the CO2/CH4 Mixture.
doi:10.1021/acs.jced.9b00503
BibTeX | RIS
2019
Austen Bernardi, Roland Faller, Dirk Reith, Karl N. Kirschner:
ACPYPE update for nonuniform 1–4 scale factors: Conversion of the GLYCAM06 force field from AMBER to GROMACS.
doi:10.1016/j.softx.2019.100241
BibTeX | RIS
Kontakt

Dirk Reith
Computational Science and Engineering , Managing Director of the TREE Institute, Faculty Advisor BRS Motorsport
Research fields
Location
Sankt Augustin
Room
B 223
Address
Grantham-Allee 20
53757 Sankt Augustin
Telephone
+49 2241 865 9678